

Cepheid, a Danaher operating company, today announced that the U.S. Food and Drug Administration (FDA) has cleared Xpert Hemorrhagic Fever Panel, a multiplex PCR test designed for the qualitative detection and identification of RNA from viruses that cause several viral hemorrhagic fevers from a single sample.

SEKISUI Diagnostics, a global medical diagnostics manufacturer, has announced the launch of a 510(k) cleared rapid test to detect COVID-19 and Flu A&B in professional healthcare settings.

bioMérieux, a world leader in the field of in vitro diagnostics, today announced that its BIOFIRE® SPOTFIRE® Vaginitis (VG) Panel has received Dual 510(k) Clearance and CLIA Waiver* from the U.S. Food and Drug Administration (FDA). This automated, multiplex polymerase chain reaction (PCR) test can simultaneously detect the most common causes of vaginitis in about 20 minutes, directly at the point of care on the BIOFIRE® SPOTFIRE® System, bringing rapid and accurate diagnostics closer to patients.
Natera, Inc. (NASDAQ: NTRA), a global leader in cell-free DNA and precision medicine, today announced that its multi-cancer early detection (MCED) assay has been granted Breakthrough Device designation (BDD) by the U.S. Food and Drug Administration (FDA). The BDD is designed for novel technologies that enable more effective diagnosis or treatment of life-threatening or irreversibly debilitating diseases or conditions.
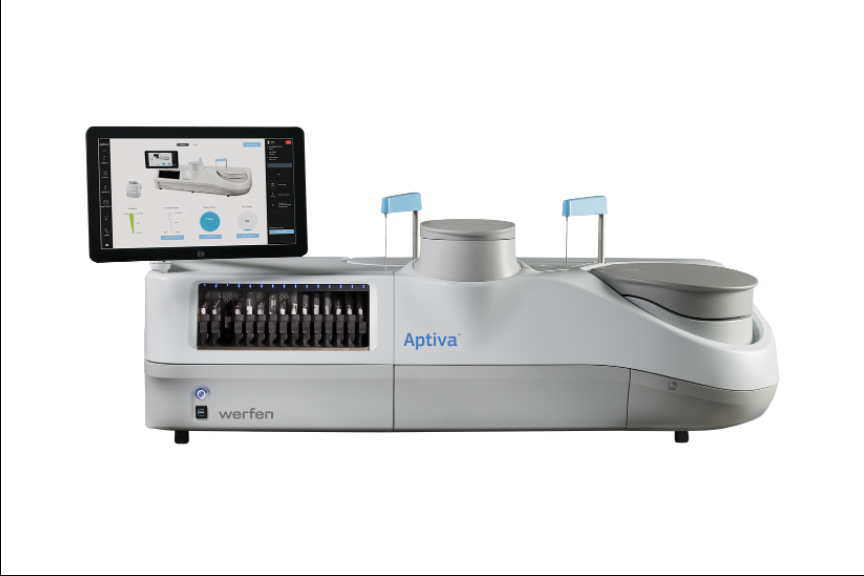
Werfen today announced that its Aptiva Antiphospholipid Syndrome (APS) Immunoglobulin A (IgA) Reagent has received both U.S. Food and Drug Administration (FDA) 510(k) clearance and CE Mark under the European Union’s In Vitro Diagnostic Medical Devices Regulation (IVDR). Together, these milestones complete the Aptiva APS testing portfolio, which now offers all three APS isotypes, IgA, IgG, and IgM, for both anti-cardiolipin (aCL) and anti-beta 2 glycoprotein 1 (aβ₂GP1) antibodies.
Teal Health, creators of the Teal WandTM, the first and most accurate FDA-authorized self-collection device for at-home HPV cervical cancer screening, today announced $22 million in Series A funding to meet accelerating consumer demand and expand access ahead of broad insurance coverage beginning in January 2027.
Pattern Bioscience, an innovator in rapid diagnostic technologies for bacterial infections, today announced that the U.S. Food and Drug Administration (FDA) has granted Breakthrough Device Designation for its Urine ID/AST Panel. The Urine ID/AST Panel is the company’s second assay to receive the designation, following the Pneumonia ID/AST Panel. It reflects Pattern’s continued focus on expansion of a menu of rapid, direct-from-specimen assays.
Guardant Health, Inc. (Nasdaq: GH), a leading precision oncology company, today announced that the U.S. Food and Drug Administration (FDA) has approved Guardant360® CDx as a companion diagnostic (CDx) to identify patients with hormone receptor (HR)-positive, HER2-negative advanced or metastatic breast cancer harboring tumors with ESR1 mutations before radiographic progression who may benefit from treatment with ETCAMAH™ (camizestrant), AstraZeneca’s next-generation oral selective estrogen receptor degrader (SERD). Guardant360 CDx is the first FDA-approved liquid biopsy CDx for longitudinal testing in breast cancer, and this approval represents a major shift from therapy selection and switching post-radiographic progression to adaptive cancer management and early interception using pre-radiographic progression circulating tumor DNA (ctDNA) testing.
Roche (SIX: RO, ROG; OTCQX: RHHBY) announced today that it has received FDA approval for expanded use of its PATHWAY® HER2 (4B5) and VENTANA® HER2 Dual ISH DNA Probe Cocktail tests. These companion diagnostics are now approved to assess HER2-positive status in metastatic gastroesophageal adenocarcinoma (GEA) patients, including those with gastric, gastroesophageal junction, and esophageal adenocarcinoma. The approval enables clinicians to identify HER2-positive GEA patients who may be eligible for HER2-targeted treatment with Jazz Pharmaceuticals' ZIIHERA® (zanidatamab-hrii). Two ZIIHERA-containing regimens are now FDA approved for the first-line treatment of adults with unresectable locally advanced or metastatic HER2-positive GEA.
Abbott (NYSE: ABT), the global healthcare leader, today announced U.S. Food and Drug Administration (FDA) authorization of its Libre Duo 10 Day system, the world's first dual glucose‑ketone sensing technology. The new biowearable is designed to alert people living with diabetes of rising ketones that can lead to a diabetic ketoacidosis (DKA) emergency, a serious health complication for people with diabetes. Libre Duo 10 Day delivers real-time visibility into glucose levels needed for daily diabetes management while also monitoring rising ketones in the background, avoiding additional mental or physical burden for people with diabetes. Abbott plans to launch Libre Duo 10 Day in the United States later this year.
✔ All (559)
✔ Press release (3)
✔ Industry news (556)
Don’t miss important updates about the show and the in vitro diagnostic industry.
Sign-up for our newsletter today.


Copyright © 2026 GL events Ruihe (Shanghai) Exhibition Co., Ltd. All Rights Reserved. ( 沪ICP备12004745号-1 )

We deliver the latest IVD news straight to your inbox. Stay in touch with CACLP News.
sign-up for our newsletter today.


To ensure our newsletter hit your inbox, make sure to add @caclp.com to your safe senders list. And, as always, feel free to contact
us with any questions and thanks again for subscribing.
Go back


